Active Litigation
Vaginal Mesh Lawsuit Facts
TorHoerman Law is actively accepting claims for the Vaginal Mesh Lawsuit.


Transvaginal mesh implants, also known as pelvic mesh implants, have been used to treat pelvic organ prolapse (POP) and stress urinary incontinence (SUI).
These conditions primarily affect pregnant women and can result in discomfort, pain, and urinary incontinence.
These conditions can lead to a range of discomforts, including pelvic pain and challenges in bladder control.
Initially, transvaginal mesh was known as a groundbreaking and permanent solution for these debilitating pelvic conditions, offering a new lease on life for many patients.
Over time, however, emerging studies and a growing body of patient testimonials have cast significant doubts on these early optimistic views.
A considerable number of women who underwent surgery involving transvaginal mesh began reporting severe complications, sometimes shortly after the procedure.
The mesh, which is designed to reinforce weakened pelvic tissues, has been found to cause issues such as mesh erosion, where the material wears through the vaginal wall, leading to excruciating pain and discomfort.
Additionally, instances of organ perforation have been documented, where the mesh punctures surrounding organs, causing severe internal damage and infection.
Many patients have reported a recurrence of their original pelvic conditions, alongside new complications such as urinary problems and discomfort during sex, negating the initial benefits of the surgery.
The severity of these complications often necessitates additional surgeries, which may not always resolve the issues and can lead to further distress and health problems.
This growing evidence of harm has led to a reevaluation of the use of transvaginal mesh implants in the medical community.
Regulatory bodies, including the U.S. Food and Drug Administration (FDA), have issued warnings and have even banned certain types of pelvic mesh products, citing safety concerns.
The issues surrounding transvaginal mesh implants have also sparked a wave of legal actions, as affected patients seek compensation for the unexpected and life-altering side effects they have endured.
A pelvic mesh implant is a medical device designed to provide additional support to weakened or damaged pelvic organs, such as the bladder, uterus, vaginal wall, or rectum, in the case of POP.
The manufacturers of these implants designed them to reinforce the pelvic floor and alleviate symptoms of organ prolapse.
However, many women in the United States have reported infectivity of such medical devices.
Complaints included pain in the abdomen and discomfort during intercourse.
Many of the correspondents claimed that they had transvaginal mesh surgery removal to alleviate the side effects.
Transvaginal mesh implants, originally designed to provide a surgical solution for pelvic conditions, have been associated with a range of serious complications.
These complications not only affect the quality of life but also pose significant health risks for patients.
Some of the most common surgical mesh complications associated with this product are:
As complications associated with transvaginal mesh implants became increasingly evident, legal action began to mount.
Patients who experienced severe side effects that often required multiple surgeries to correct sought justice through lawsuits against manufacturers.
Many of these cases alleged that manufacturers failed to warn patients and medical providers adequately about the potential risks.
Some of the involved manufacturers named in Vaginal Mesh Lawsuits include:
Because of the upsurge in lawsuits, the court consolidated similar cases into multidistrict litigation (MDL).
These lawsuits also prompted the FDA to look closer at the safety and effectiveness of transvaginal mesh implants.
Today, most MDLs and lawsuits against manufacturers have been settled, and experts believe that the settlements and verdicts could collectively reach up to $11 billion.
Even though most surgical mesh lawsuits have been resolved, these cases can still be filed in relevant courts.
If you or a loved one have suffered injuries due to defective vaginal mesh, you may be eligible to file a vaginal mesh lawsuit.
Contact TorHoerman Law for a free consultation, or use the chatbot on this page to find out if you qualify for the vaginal mesh lawsuit instantly.
The timeline of FDA recalls concerning transvaginal mesh implants is a testament to the intricate and evolving nature of medical device regulation and patient safety.
Initially introduced as a revolutionary solution for pelvic organ prolapse (POP) and stress urinary incontinence (SUI), transvaginal mesh implants soon became a subject of concern, leading to a series of regulatory actions and recalls by the U.S. Food and Drug Administration (FDA).
This journey began in 2008, marking the start of an extensive period of scrutiny and reassessment that spanned over a decade.
In response to mounting reports of complications and adverse events associated with these implants, the FDA issued its first public health notification.
This initial step was a significant move by the regulatory agency, acknowledging the potential risks and signaling the need for closer observation.
As patient complaints and clinical evidence continued to accumulate, the FDA’s stance on transvaginal mesh implants evolved.
The agency’s efforts included rigorous analysis of clinical data, consultations with healthcare professionals and patients, and careful monitoring of post-market device performance.
These efforts culminated in a series of recalls and safety communications, each marking a critical point in the ongoing evaluation of the safety and efficacy of these devices.
Throughout this period, the FDA’s actions reflected a growing awareness of the complexities associated with transvaginal mesh implants.
The recalls were not just administrative measures — they represented an ongoing commitment to patient safety and the need for a balanced approach in the use of medical devices.
In October 2008, the FDA released a public health notification titled “Serious Complications Associated with Transvaginal Placement of Surgical Mesh in Repair of Pelvic Organ Prolapse and Stress Urinary Incontinence.”
This notification raised concerns about the severe complications linked to the placement of the transvaginal mesh device to treat organ prolapse and stress urinary incontinence.
The FDA’s communication was a significant turning point in the history of transvaginal mesh implants.
It marked the beginning of increased scrutiny and awareness of the risks associated with these devices.
The FDA further addressed the concerns surrounding transvaginal mesh implants in 2011.
The agency issued an updated safety communication about the complications associated with these devices, specifically for treating POP.
This communication highlighted the increasing evidence of complications and urged healthcare providers to be cautious when considering surgical mesh.
The FDA’s involvement in investigating these issues intensified.
In 2011, the FDA convened the Obstetrics and Gynecology Devices Panel of the Medical Devices Advisory Committee.
This meeting aimed to discuss the safety and effectiveness of surgical mesh used for both SUI and POP.
During the meeting, the panel recommended reclassifying the surgical mesh used for transvaginal repair of POP from Class II (low-to-moderate risk devices) to Class III (high-risk devices).
This change would make the regulatory requirements more stringent for manufacturers.
In response to mounting concerns, the FDA ordered the 522 Postmarket Surveillance Studies Program, often referred to as “522 studies,” to be conducted by manufacturers of transvaginal surgical mesh devices.
These studies were initiated to address specific safety and effectiveness concerns related to mini-sling devices for SUI and surgical mesh used for transvaginal repair of POP.
By February 17, 2013, the FDA had issued 95 postmarket study orders to 34 surgical mesh manufacturers for POP and 14 postmarket study orders to seven manufacturers of mini-slings for SUI.
These studies aimed to provide a clearer understanding of the safety and effectiveness profiles of these devices.
In 2013, the FDA updated its information about stress urinary incontinence (SUI) on its website.
This update included the FDA’s current thinking on the use of surgical mesh for the repair of SUI.
The update was based on an analysis of adverse events reported to the FDA, findings in the scientific literature, and input received from the 2011 meeting of the Obstetrics and Gynecology Devices Panel of the Medical Devices Advisory Committee.
The FDA aimed to provide patients with more comprehensive information about the risks and benefits of using surgical mesh for SUI.
In 2014, the federal agency presented two orders related to surgical mesh for transvaginal pelvic organ prolapse (POP) repair.
These orders were introduced to address the risks associated with transvaginal mesh devices more effectively:
The proposed orders issued in 2014 were finalized by the FDA in 2016.
Because of these orders, the surgical mesh for POP was reclassified as Class III, marking it as a high-risk device.
This change required manufacturers to submit PMA applications by July 5, 2018, for any surgical mesh intended for transvaginal POP repair.
The FDA’s actions led to all manufacturers ceasing the marketing of surgical mesh intended to treat pelvic organ prolapse or any form of posterior compartment prolapse.
This reclassification was a significant development in addressing the risks associated with transvaginal mesh implants.
Following the reclassification of the product and its changes in its intended use, the agency called the attention of the last mesh surgical product manufacturer.
The agency demanded the manufacturer halt its production and distribution efforts.
Subsequently, the company voluntarily withdrew its product from the market.
This action represented a further reduction in the availability of transvaginal mesh implants, signaling the FDA’s commitment to addressing the risks associated with these devices.
The FDA reconvened the Obstetrics and Gynecology Devices Panel of the Medical Devices Advisory Committee.
In this meeting, the agency sought expert insights in evaluating the safety and effectiveness of surgical mesh for transvaginal repair of prolapse.
The meeting concluded, and the committee decided that for the vaginal mesh implants to be considered effective, they must surpass the native tissue repair rate of 36 months.
This criterion meant that the mesh should yield a more efficient healing capability than the body’s natural way of healing.
After reclassifying surgical mesh for transvaginal repair of pelvic organ prolapse into Class III, the FDA initiated a review of two Premarket Approval Applications (PMA) submitted by Boston Scientific for its devices, the Uphold LITE Vaginal Support System and the Xenform Soft Tissue Repair System.
Additionally, the agency reviewed the PMA application from Coloplast’s team for their Restorelle DirectFix Anterior.
With the new criterion in effect, the FDA denied the premarket applications of these manufacturers because of their inefficiency in treating POP.
The agency found that the benefits of using the product did not outweigh its potential risks.
Consequently, the FDA ordered the two manufacturers of these mesh surgical products to cease selling and distributing their devices.
Both companies voluntarily withdrew their products from the market.
Despite the agency’s effective ban on Boston Scientific’s Uphold Lite and Xenform, the FDA required the manufacturer to continue following up with patients who were already enrolled in their postmarket surveillance studies (“522 studies”).
These studies evaluated the effectiveness and safety of transvaginal mesh for POP compared to the body’s healing process without the mesh.
The two 522 studies concluded with a negative outcome.
The product only produced remedial effects after 36 months, similar to native healing.
Including the 36-month follow-up data, the FDA found that the manufacturer’s mesh product is less effective than the natural healing process.
Similar to Boston Scientific, the agency demanded Coloplast to maintain communication regarding the patients already participating in the Coloplast 522 postmarket study.
Likewise, the manufacturer’s Direct Fix A and Direct Fix P produced similar results with Boston Scientific’s Uphold Lite and Xenform.
The study showed that Coloplast transvaginal POP mesh had identical effectiveness and safety outcomes to native tissue repair at 36 months.
Because of its inefficiency in treating organ prolapse, the FDA maintained the ban on transvaginal mesh implants.
The FDA’s commitment to ensuring the safety of these transvaginal surgical mesh devices is shown through regulatory guidance.
Currently, the agency is executing progressive efforts to mitigate its risks in several ways:
The FDA has yet to conduct a comprehensive analysis of transvaginal mesh products from all involved manufacturers.
If you’re suffering from organ prolapse or stress urinary incontinence after pregnancy or menopause, it’s best to speak with your gynecologists about proven treatment and alternatives from these devices
The recalls and regulatory changes surrounding transvaginal mesh implants have significant implications for patients.
These actions are ultimately aimed at improving patient safety and ensuring that the risks associated with surgical mesh are thoroughly assessed and addressed.
For patients who have received transvaginal mesh implants, it’s essential to be aware of the evolving regulatory landscape and to stay informed about the latest developments.
If you have experienced complications or discomfort related to a mesh implant, it’s advisable to consult with your healthcare provider to discuss your options and seek appropriate medical care.
Although most MDLs have been settled, you may still be able to file a lawsuit.
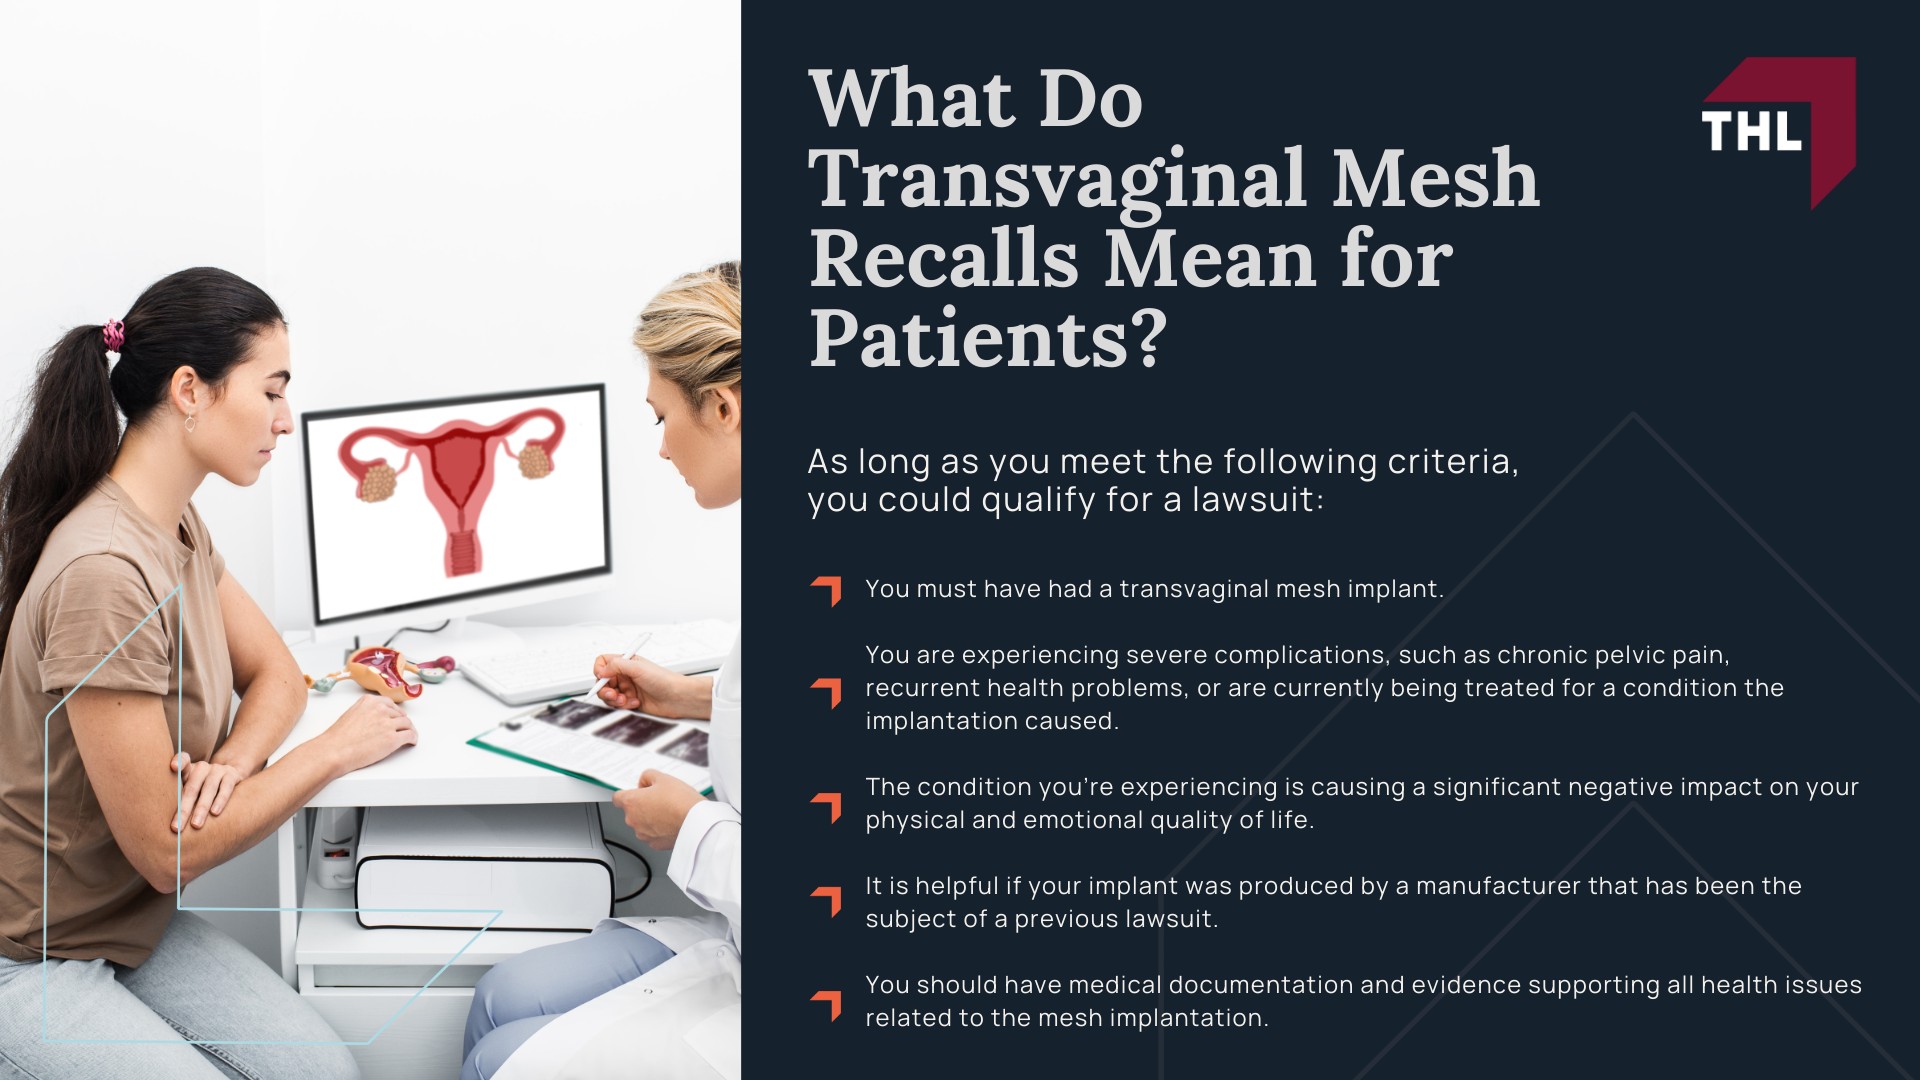
As long as you meet the following criteria, you could qualify for a lawsuit:
Aside from your tangible documents, hiring the right legal team increases the chances of success in your case.
At TorHoerman Law, our team has experience handling cases related to medical products, and the transvaginal surgical mesh is no exception.
We can help you claim compensation for your damages, such as:
The FDA’s evolving approach to regulation, along with the legal actions patients have taken, has resulted in greater awareness and enhanced patient safety.
Patients who have experienced complications related to transvaginal mesh should stay informed and consider seeking legal representation to address their concerns.
If you believe you have a case related to transvaginal mesh complications, our legal team can help.
TorHoerman Law is a prominent law firm with a strong track record in handling issues related to defective medical products.
Contact us now or use our chatbot for a quick and free case evaluation.


